Home / About LSDs / Gaucher Disease
Gaucher Disease: Symptoms, Treatment and Support in India
Gaucher disease is a rare inherited disorder in which a fatty substance builds up in the spleen, liver and bones. Effective treatment is available, and LSDSS has supported families across India since 2010.
Key facts
Cause
Deficiency of the enzyme glucocerebrosidase (GBA1 gene)
Inheritance
Autosomal recessive
Mainly affects
Spleen, liver, bone marrow and bones
Treatment
Enzyme replacement therapy (ERT) and substrate reduction therapy (SRT)
LSDSS community
Over 200 registered Gaucher families across India
On this page:
Overview
What is Gaucher disease?
Gaucher disease is a rare genetic disorder in which the body cannot properly break down a fatty substance called glucocerebroside. This happens because of a deficiency of the enzyme glucocerebrosidase.
The fatty substance builds up inside cells called Gaucher cells, mainly in the spleen, liver and bone marrow. This can cause an enlarged spleen and liver, anaemia, easy bruising, and bone pain and fractures.
In India, Gaucher disease is often diagnosed late because its symptoms can look like those of more common conditions.
Video: Gaucher disease explained (credit: Osmosis)
Gaucher disease in India
Gaucher disease affects an estimated 1 in 40,000 to 60,000 people worldwide. In India, the true number is hard to estimate because many patients are undiagnosed or misdiagnosed.
Growing awareness and better diagnostic facilities are helping to identify more patients. Some communities see more cases because of consanguinity (marriage between close relatives), which raises the risk of inherited conditions like Gaucher disease. This is why genetic awareness and early screening matter.
Did you know?
LSDSS supports over 200 registered Gaucher families across India.
Early diagnosis and timely treatment make a major difference to how patients do.
Types of Gaucher disease
Doctors describe three main types, based on whether the brain and nervous system are affected.
Type 1 (non-neuronopathic)
The most common form. It mainly affects the spleen, liver, blood and bones, and does not affect the brain. Symptoms can appear at any age, from childhood to adulthood, and respond well to treatment.
Type 2 (acute neuronopathic)
A rare, severe form that begins in infancy. It affects the brain and nervous system as well as the organs, and progresses quickly. Current treatments cannot reverse its effects on the brain.
Type 3 (chronic neuronopathic)
Usually starts in childhood or adolescence. It affects the organs and, more slowly, the brain, for example eye movements and coordination. Treatment helps the organ and blood symptoms.
Signs and symptoms
Symptoms of Gaucher disease
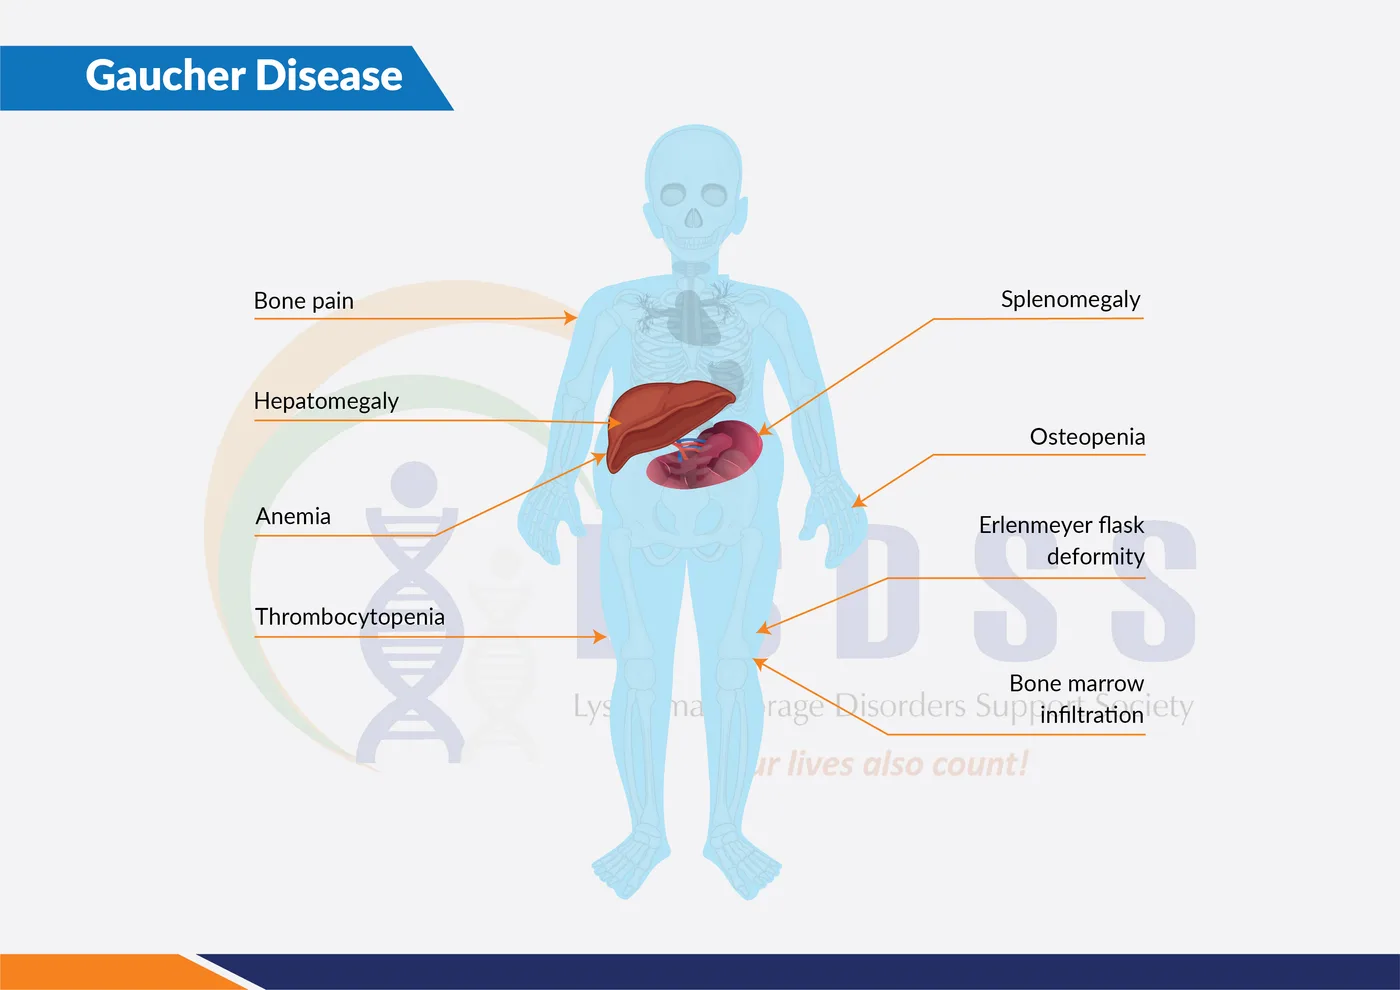
Symptoms vary from person to person, even within the same family. Common signs include:
- Enlarged spleen, causing a swollen abdomen
- Enlarged liver
- Tiredness and weakness due to anaemia
- Easy bruising or bleeding due to low platelets
- Bone pain, bone crises and frequent fractures
- Delayed growth or puberty in children
When to see a doctor
Consult a doctor, ideally a geneticist or haematologist, if you or your child has:
- A persistently enlarged spleen
- Unexplained anaemia or low platelets
- Frequent bone pain or fractures
- Easy bruising or nosebleeds
These signs are often mistaken for other conditions in India, so ask about Gaucher disease if they persist.
Causes
What causes Gaucher disease?
Gaucher disease is caused by changes (mutations) in the GBA1 gene. This gene gives the body instructions to make the enzyme glucocerebrosidase.
When the enzyme is missing or does not work properly, glucocerebroside builds up in organs and tissues and causes the symptoms of the disease.
How it is inherited
- It is inherited in an autosomal recessive pattern.
- Parents of an affected child are usually healthy carriers of one faulty copy of the gene.
- When both parents are carriers, each pregnancy has a 1 in 4 (25%) chance of an affected child.
- Genetic counselling and carrier testing can help families understand their risk and plan.
How is Gaucher disease diagnosed?
Early testing at a specialised centre leads to timely treatment and better outcomes.
01
Enzyme test
A blood test measures glucocerebrosidase activity. Low activity points to Gaucher disease.
02
Genetic test
Confirms changes in the GBA1 gene and helps with testing other family members.
03
Blood tests
Check for anaemia, low platelets and other markers of the disease.
04
Imaging
MRI or CT scans measure the spleen and liver and check the health of the bones.
Treatment for Gaucher disease
There is no cure yet, but effective treatments can greatly improve quality of life.
Enzyme replacement therapy (ERT)
The standard treatment. The missing enzyme is given as an intravenous infusion, usually every two weeks, for life. It reduces liver and spleen size, improves blood counts and reduces bone problems. It works well for Type 1 but does not improve neurological symptoms in Types 2 and 3.
Medicines include
- Imiglucerase (Cerezyme®)
- Velaglucerase alfa (VPRIV®)
- Taliglucerase alfa (Elelyso®)
Substrate reduction therapy (SRT)
An oral (tablet) option for some adults with Type 1 Gaucher disease. Instead of replacing the enzyme, it reduces how much of the fatty substance the body makes.
Medicines include
- Eliglustat (Cerdelga®)
- Miglustat (Zavesca®)
Supportive and long-term care
Depending on their symptoms, patients may also need:
This can include
- Blood transfusions for severe anaemia
- Pain management for bone disease
- Regular monitoring with blood tests and MRI
- Physiotherapy and orthopaedic care
Research into gene therapy, which could one day offer a one-time treatment for Gaucher disease, is under way.
Clinical trials and studies in India
Studies of Gaucher disease that are recruiting, or about to recruit, in India. This list updates automatically every day.
- Study to Evaluate the Efficacy and Safety of Nizubaglustat (AZ-3102) in Patients With Gaucher Disease Type 3 (GD3)Not yet recruitingPhase 2 treatment trial · Azafaros B.V.Sites in India: Jaipur, New Delhi
- International Collaborative Gaucher Group (ICGG) Gaucher Disease Registry & Pregnancy Sub-registryRecruitingObservational study or registry · Genzyme, a Sanofi CompanySites in India: Ahmedabad, Jaipur, Kolkata, Mumbai, Secunderabad, Vellore
Last checked 13 November 2026 on ClinicalTrials.gov. Always talk to your doctor before joining a study.
Accessing treatment in India
Getting treatment and financial support
Treatment for Gaucher disease is expensive, costing from several lakhs to a few crore rupees a year, lifelong. These are the main sources of help.
Government support (NPRD)
Under the National Policy for Rare Diseases, eligible patients can receive financial assistance of up to ₹50 lakh for treatment at a designated Centre of Excellence (COE).
- Visit your nearest COE with all medical reports
- Assistance is subject to approval
- The ₹50 lakh limit may not cover lifelong ERT
Other sources of help
- State government schemes
- Corporate social responsibility (CSR) funding
- Crowdfunding through the government rare disease portal
- Charitable access programmes (limited places)
How LSDSS helps
- Connects patients with COEs and specialists
- Guides families through funding applications
- Shares updates on support programmes
- Brings patients and caregivers together
Frequently asked questions
Answers to common questions about Gaucher disease.
Still have a question?
A rare inherited disorder in which a deficiency of the enzyme glucocerebrosidase causes a fatty substance to build up in the spleen, liver, bone marrow and bones.
The most common signs are an enlarged spleen (a swollen abdomen), an enlarged liver, tiredness and weakness from anaemia, easy bruising or bleeding from low platelets, and bone pain or fractures. Children may grow or reach puberty late. Types 2 and 3 also affect the brain and nervous system.
It depends on the type. With treatment, most people with Type 1 Gaucher disease live full lives. Type 2 is severe and usually fatal in early childhood. Type 3 shortens life expectancy, but treatment helps many patients live into adulthood.
Not yet. Enzyme replacement therapy and substrate reduction therapy manage the disease effectively. Research into gene therapy, which could offer a one-time treatment, is under way.
It is inherited. A child is affected when both parents pass on a faulty copy of the GBA1 gene. Parents who are carriers usually have no symptoms.
According to the International Collaborative Gaucher Group (ICGG) Gaucher Registry, average life expectancy at birth for people with Type 1 Gaucher disease is about 68 years (about 64 years for those whose spleen was removed and 72 years for those whose spleen was not), compared with about 77 years in a similar reference population. Type 2 usually results in death within the first few years of life. Type 3 shortens life expectancy, but treatment helps some patients with milder neurological involvement live into their 50s.
Because it is genetic, it cannot be prevented in the usual sense. Genetic counselling, carrier testing and prenatal testing can help families with a history of Gaucher disease make informed choices.
First, visit your nearest Centre of Excellence (COE) with all medical reports, to be evaluated and enrolled on the Government of India’s rare disease portal for crowdfunding and CSR donations. Second, register with LSDSS for support and the latest information. You can also ask to join the International Gaucher Alliance (IGA) waiting list for charitable treatment by filling in the IGA online form. Please also let us know at contactus@lsdssindia.org once you have applied. Charitable programmes currently have very limited places.
Other lysosomal storage disorders
LSDSS supports patients and families living with all lysosomal storage disorders.
A fatty substance builds up in nerve cells of the brain and spinal cord.
Sulfatides build up and damage the protective covering of nerves.
A fatty substance builds up in nerve cells and, in babies, in other organs and bones.
Waste material builds up in nerve cells, causing seizures and loss of vision.
Living with Gaucher disease?
Register with LSDSS for treatment guidance, support and the latest updates. You are not alone.
This page is for general information and awareness only. It does not replace advice from a qualified doctor. Last reviewed: November 2026.